
Virtual ScreeningPharmaceutical InformationUpdated on Jul 30, 2019 View more like this | Visit SHIRLEY, NY | Contact BOCSCI INC |
Advantages of Our Services:
Integrated in silico and experimental team of multi-disciplinary experts in chemistry, biology and computing sciences
Ligand-based and structure-based virtual screening
Super high-performance computer cluster with total 60 blades and 720 cores
Compound database containing over 10 million purchasable compounds
Compound database compliant with predefined filtering rules
Molecular docking, molecular dynamics simulations, free energy calculations
3D pharmacophore model building
Affinity prediction, fragment based approaches, handling of protein flexibility
Consideration of water and solvation effects
Virtual Screening
Structure-based virtual screening
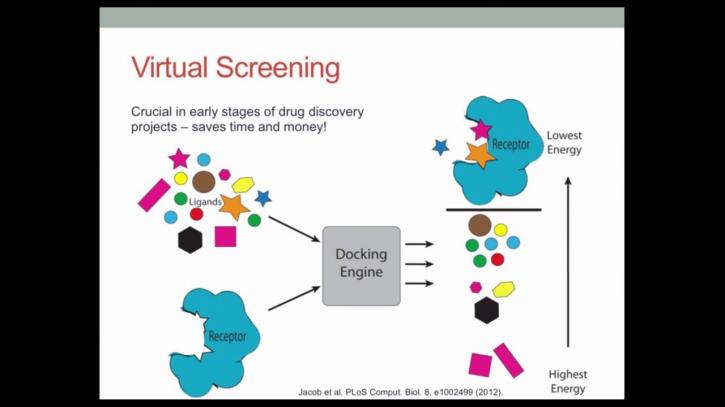
The general scheme of a Structure-Based Virtual Screening (SBVS) strategy starts with processing the 3D target structural information of pharmaceutical protein interested. The target structure can be derived from experimental data (X-ray, NMR or neutron scattering spectroscopy), homology modeling, or from Molecular Dynamics (MD) simulations. The identification of ligand binding sites on biological targets is incredibly important. Our expertise evaluates the druggability of the receptor, the choice of binding site, the selection of the most relevant protein structure, incorporating receptor flexibility, suitable assignment of protonation states, and consideration of water molecules in a binding site. Our highly experienced drug design professionals carefully choose the compound library to be screened in the virtual screening (VS) exercise according to the target in question, and preprocess libraries to assign the proper stereochemistry, tautomeric, and protonation states.
Integrated in silico and experimental team of multi-disciplinary experts in chemistry, biology and computing sciences
Ligand-based and structure-based virtual screening
Super high-performance computer cluster with total 60 blades and 720 cores
Compound database containing over 10 million purchasable compounds
Compound database compliant with predefined filtering rules
Molecular docking, molecular dynamics simulations, free energy calculations
3D pharmacophore model building
Affinity prediction, fragment based approaches, handling of protein flexibility
Consideration of water and solvation effects
Virtual Screening
Structure-based virtual screening
The general scheme of a Structure-Based Virtual Screening (SBVS) strategy starts with processing the 3D target structural information of pharmaceutical protein interested. The target structure can be derived from experimental data (X-ray, NMR or neutron scattering spectroscopy), homology modeling, or from Molecular Dynamics (MD) simulations. The identification of ligand binding sites on biological targets is incredibly important. Our expertise evaluates the druggability of the receptor, the choice of binding site, the selection of the most relevant protein structure, incorporating receptor flexibility, suitable assignment of protonation states, and consideration of water molecules in a binding site. Our highly experienced drug design professionals carefully choose the compound library to be screened in the virtual screening (VS) exercise according to the target in question, and preprocess libraries to assign the proper stereochemistry, tautomeric, and protonation states.
